


*****The following content was written by the author(s) of the paper.*****
A research team has identified the transcription factor C/EBPγ as a novel regulator that simultaneously promotes epithelial-mesenchymal transition (EMT) and DNA double-strand break repair in lung adenocarcinoma cells. The findings provide new insights into how cancer cells acquire aggressive features and resistance to anticancer treatments.
EMT is a reversible cellular program in which epithelial cancer cells acquire mesenchymal characteristics, enhancing their mobility, invasiveness, and adaptability. This process is closely associated with tumor progression, metastasis, and resistance to therapy. Although several major EMT-inducing transcription factors have been identified, less is known about factors that connect EMT with other malignant properties that help cancer cells survive treatment. To identify such regulators, the researchers employed an epigenomic approach that focused on broad domains of histone H3 lysine 4 trimethylation (H3K4me3), a chromatin signature associated with genes that define cellular identity. By examining genes whose H3K4me3 domains expanded during TGF-β-induced EMT, the team identified C/EBPγ as a candidate EMT regulator.
Further experiments demonstrated that introducing C/EBPγ into lung adenocarcinoma cells induced hallmark EMT features, including elongated mesenchymal-like morphology, reduced E-cadherin expression, increased expression of mesenchymal markers, and enhanced migratory capacity. In contrast, depletion of endogenous C/EBPγ suppressed EMT-associated gene expression and impaired EMT progression. The researchers discovered that C/EBPγ operates through an unusual mechanism. Unlike many transcription factors that require direct DNA binding, C/EBPγ-induced EMT depended on its leucine zipper domain but not on its DNA-binding domain. This finding suggested that interactions with other proteins are critical for its biological activity. Proteomic analyses identified several C/EBPγ-interacting proteins, including C/EBPβ and the DNA repair factors XRCC5 and XRCC6. Additional studies revealed that C/EBPβ functions as an EMT suppressor in lung adenocarcinoma cells, whereas C/EBPγ promotes EMT by antagonizing C/EBPβ activity through leucine zipper-dependent interactions.
The study also uncovered a second and independent role for C/EBPγ in DNA repair. XRCC5 and XRCC6 are core components of the non-homologous end joining (NHEJ) pathway, a major mechanism used by cells to repair DNA double-strand breaks. The investigators found that C/EBPγ physically associates with these proteins and promotes NHEJ activity. C/EBPγ accelerated recruitment of XRCC6 to damaged DNA, reduced accumulation of DNA damage markers following etoposide treatment, and enhanced overall repair efficiency. Importantly, these molecular effects translated into increased resistance to therapy. Lung adenocarcinoma cells expressing C/EBPγ survived DNA-damaging chemotherapy more effectively than control cells. In mouse xenograft models, tumors expressing C/EBPγ remained significantly less sensitive to etoposide treatment, whereas disruption of the leucine zipper domain abolished this protective effect.
The findings indicate that C/EBPγ acts as a molecular hub connecting EMT-associated cellular reprogramming with enhanced DNA repair capacity. By coordinating these two mechanisms, C/EBPγ may help tumor cells adapt to therapeutic stress and acquire treatment resistance. The researchers conclude that targeting C/EBPγ or its protein-interaction interfaces could represent a new strategy for improving the effectiveness of DNA-damaging therapies and overcoming therapy resistance in lung adenocarcinoma.
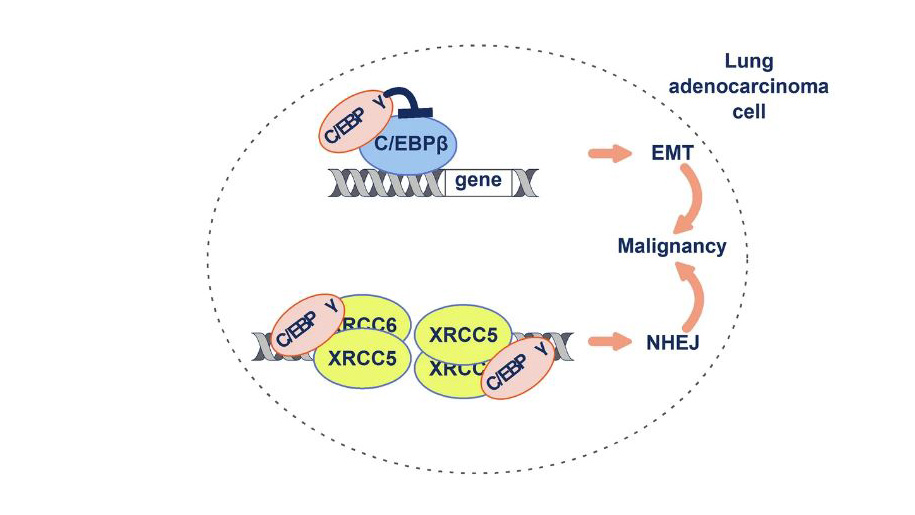
Figure: Mechanistic model of C/EBPγ-mediated EMT, DNA repair, and therapy resistance in lung adenocarcinoma.
Proposed model illustrating how C/EBPγ promotes epithelial-mesenchymal transition (EMT) through antagonism of C/EBPβ while enhancing DNA double-strand break repair through interactions with XRCC5/XRCC6. These dual functions increase survival of lung adenocarcinoma cells under genotoxic stress and may contribute to therapy resistance.
Adapted from Terashima, M. et al.Cell Death Discovery, 2026, © The Authors, CC BY 4.0.
Click here to see the press release.【Japanese only】
Journal : Cell Death Discovery
Researcher Information : Takeshi Suzuki
Related Information
Cancer Research Institute, Kanazawa University
Graduate School of Natural Science and Technology, Kanazawa University
 PAGE TOP